
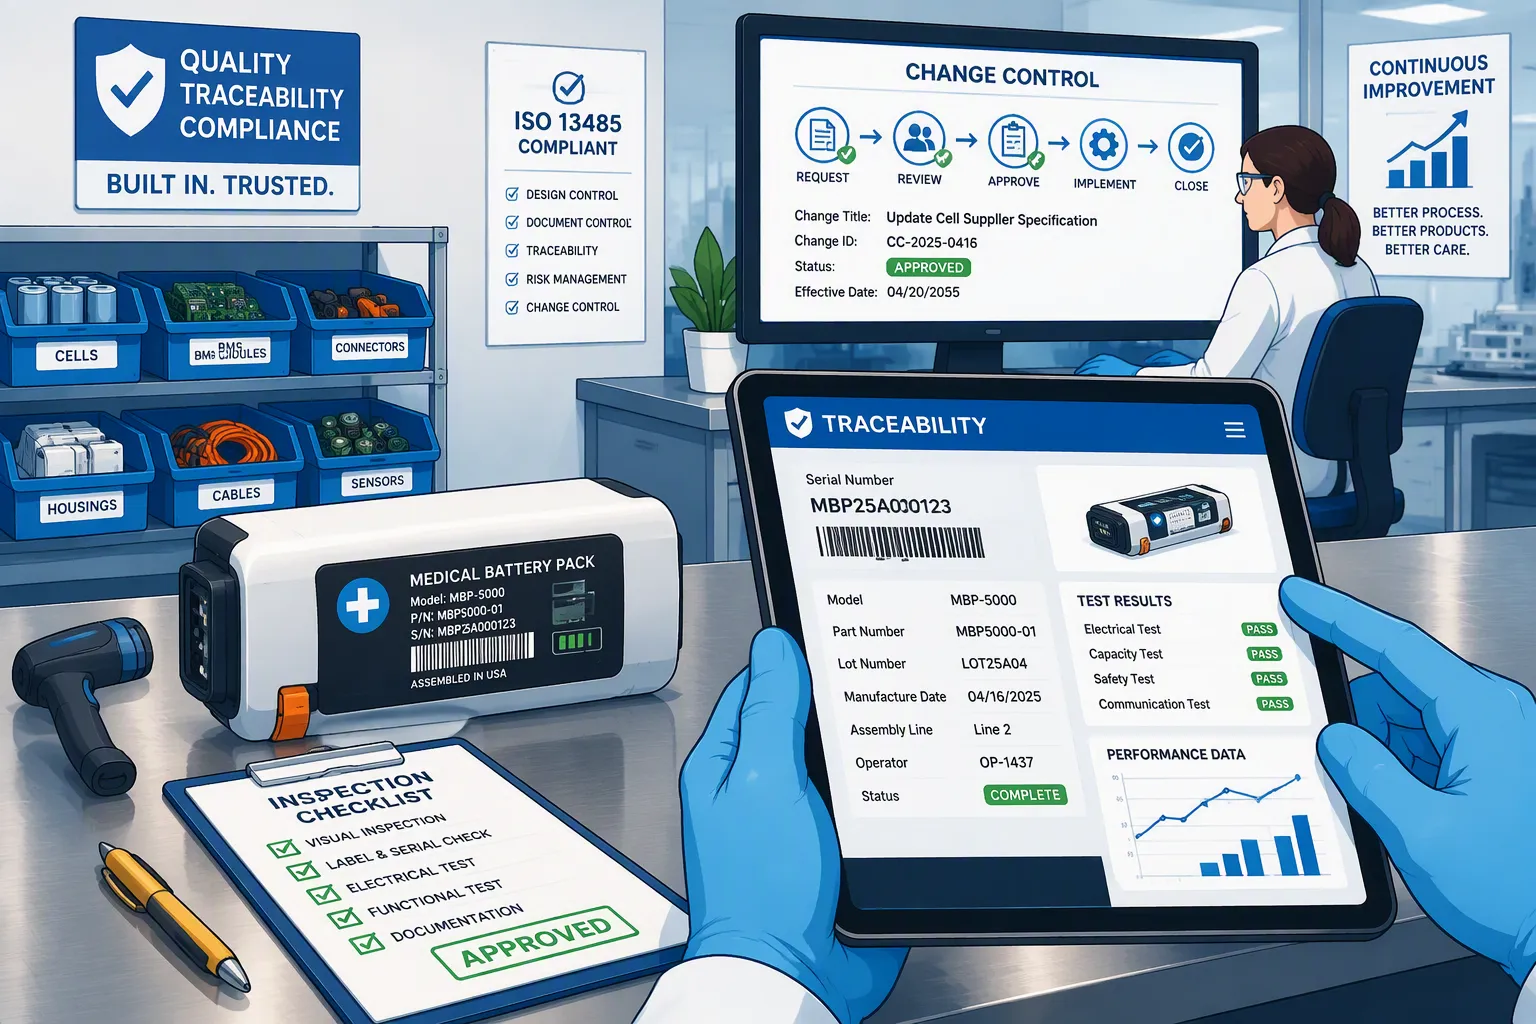
When a medical device develops a battery-related fault, engineers need to identify the affected cell lots, pack configurations, and BMS revisions. Reliable records help establish the investigation scope; they do not, by themselves, prove which component caused the failure.
For medical battery packs, traceability and change control should connect the approved design to the product actually manufactured and supplied. This article outlines a practical approach for pack manufacturers and medical device OEMs working within an ISO 13485 quality management framework.
Quick Answer: Link each pack identifier to its cell lot, approved bill of materials, BMS hardware and firmware revisions, production records, and release results. Define traceability depth and supplier responsibilities according to applicable requirements and risk. Evaluate changes before implementation, document verification and any necessary validation, and record exactly when an approved revision enters production.
Key Takeaways
Distinguish cell supplier records, pack manufacturing records, and finished-device records.
ISO 13485 does not prescribe a universal Data Matrix code or MES platform for every battery cell.
Control cell substitutions, BMS firmware, protection settings, and critical manufacturing changes.
Use design risk analysis and process risk analysis for their respective purposes.
Test record retrieval and correct missing links before a real field investigation.
Part1: Define the Traceability Scope
1.1 Separate Quality-System Requirements from Product Approval
ISO 13485:2016 concerns quality management systems for medical devices and related services. Certification of a quality system is not approval of an individual battery design, and it does not replace device-level safety and performance evaluation.
The traceability procedure should define its scope and records in line with applicable regulatory requirements. Do not apply requirements specific to implantable devices indiscriminately to every external medical battery pack. Customer agreements and the device risk assessment can require additional controls.
FDA’s Quality Management System Regulation (QMSR) became effective on February 2, 2026 and incorporates ISO 13485:2016 by reference, with additional FDA requirements. Determine applicability according to the company’s role and the product supplied; a component supplier should not assume that its obligations are identical to those of a finished-device manufacturer.
1.2 Allocate Responsibilities Across the Supply Chain
The cell manufacturer normally maintains its internal material and process genealogy. The pack manufacturer links purchased cell lots to pack production and release records. The medical device OEM connects the battery configuration to the finished device and its distribution records.
Access to separator or electrolyte batch information may require supplier cooperation. It is not necessary to imply that every pack factory directly measures or stores all upstream raw-material data. Agree which records are retained by each party, how long they remain available, and how they can be retrieved during an investigation.
Part2: Build a Usable Pack History
2.1 Connect Incoming Materials to Released Packs
The following is an engineering checklist, not a universal list mandated for every battery product.
Record | Practical information to retain |
|---|---|
Cell identity | Manufacturer, model, supplier lot, receiving lot, quantity and acceptance status |
Pack identity | Pack serial number or controlled batch identifier, model and BOM revision |
BMS configuration | Board revision, firmware version and controlled protection/configuration settings |
Assembly history | Work order, relevant process revision, inspections and approved rework |
Release evidence | Test procedure version, results, acceptance decision and authorization |
Delivery link | Customer, shipment record and identifiers supplied for OEM traceability |
For custom lithium battery packs, serial-level pack records often make service and investigation easier. Individual cell serialization can add detail where available and justified. However, controlled lot segregation and assembly records can also maintain genealogy. A cell without an individual Data Matrix code is not automatically untraceable.
2.2 Control Rework, Firmware Updates and Lot Mixing
Record both the original and replacement components when rework changes a released configuration. Preserve firmware update history rather than overwriting the previous version. If more than one approved cell lot enters a pack batch, record the actual allocation and prevent accidental mixing with unapproved material.
A spreadsheet, database or MES must be suitable for its intended use. Define permissions, backups, change history and retrieval controls. Assess validation needs for software used in the quality system according to its application and risk; purchasing an MES does not automatically establish compliant records.
2.3 Test Forward and Backward Retrieval
Start with a cell lot and identify the associated packs and shipments. Then select a released pack and retrieve its component and configuration history. Include stock, work in progress, reworked units and returned products where relevant.
Set exercise frequency and response targets through the quality procedure and customer requirements. An annual exercise or an hours-based retrieval target may be useful, but neither should be presented as a universal ISO 13485 rule. Record gaps, corrective actions and follow-up checks.
Part3: Evaluate Changes Before Production Release
3.1 Establish Supplier Change Notification
Define notification expectations for changes that could affect purchased-product requirements. Relevant examples include cell model or construction, manufacturing location, material formulation, critical dimensions and discontinued components. Specify the information required, review contacts and implementation timing in the supplier agreement.
A supplier’s assurance of equivalent capacity is insufficient to approve a replacement cell. Review voltage limits, impedance, pulse capability, temperature limits, dimensions, aging behavior and supporting safety documentation. Keep supplier approval separate from approval of the exact part and revision.
3.2 Match the Review to the Change
Change | Potential impact | Example evaluation |
|---|---|---|
Cell model substitution | Runtime, voltage sag, charging and thermal behavior | Electrical comparison, representative load tests and safety-document review |
BMS firmware or settings | Cutoffs, alarms, SOC reporting and recovery behavior | Software review, regression tests and fault-response checks |
MOSFET, fuse or connector | Heating, interruption capacity and voltage loss | Worst-case current, thermal and protection coordination checks |
Welding process | Joint strength, resistance and consistency | Process risk review and verification or validation as appropriate |
Enclosure or insulation | Mechanical protection, spacing and heat transfer | Fit, mechanical, insulation and thermal evaluation |
Use design risk analysis for changes to the battery architecture and protective functions. PFMEA addresses manufacturing process failure modes; it is not a substitute for device-level risk management. Neither a low risk-priority number nor a supplier certificate alone establishes acceptability.
3.3 Control Implementation and Existing Inventory
Document the change rationale, affected requirements, risk assessment, verification results, necessary validation, approvals and effective serial number or lot. Decide how to handle existing stock and units already delivered. Coordinate OEM approval and regulatory or certification impact reviews where applicable.
Not every change requires complete retesting or a new certification. The evaluation must justify the scope. Changing chemistry, voltage window or safety-critical behavior can warrant substantial redesign and system-level validation.
Part4: Keep the System Effective
4.1 Use Risk-Based Supplier and Process Controls
Supplier evaluation should reflect component criticality and supplier performance. Document reviews, sample qualification and audits can all contribute. Do not confuse certification audit stages with a mandatory two-stage audit of every supplier.
Assess assembly processes whose results cannot be fully verified by later inspection. Define suitable validation and monitoring where required. The exact controls depend on the process; a universal claim that every welding operation requires the same IQ/OQ/PQ package is too broad.
4.2 Connect Investigations to CAPA
Traceability narrows an investigation; test results and root-cause analysis determine the appropriate action. Update supplier controls, design records, work instructions or training when justified. Verify effectiveness before closure.
Choose CAPA metrics and deadlines according to risk and the quality system. There is no universal battery-specific requirement for an 85% closure rate, a 60-120-day completion window or a fixed financial saving. Record-retention periods should reflect applicable requirements, product lifetime and contractual obligations.
A practical next step is to review one released pack from incoming cell receipt through customer shipment, then walk one recent change through approval and implementation. For project-specific support, discuss the required documentation and configuration controls through Large Power’s custom battery consultation.
FAQ
Does every cell need a unique serial number?
Not universally. Define identification granularity from applicable requirements and risk. Lot-based cell records linked to identifiable packs can support traceability when segregation and assembly records are reliable.
Must a pack manufacturer retain electrolyte and separator batch records?
Those records are usually maintained upstream by the cell manufacturer. Define the required access and retention arrangements with suppliers rather than assuming every pack factory holds the complete raw-material genealogy.
Does a BMS firmware update require change control?
Yes. Evaluate its impact on requirements, protection behavior, communications and device integration. Document testing and approval, and retain the version history for affected packs.
Does every replacement cell require full recertification?
No universal answer applies. Review the change against the approved design, existing reports and applicable requirements, then agree the necessary verification, validation and certification actions with the responsible parties.
Can traceability prevent all recalls?
No. It helps identify potentially affected products and supports investigation and corrective action. It does not replace design validation, manufacturing controls or an evidence-based field safety assessment.


